Common EU GMP Compliance Challenges for Pharmaceutical Manufacturers and How to Address Them
Maintaining consistent pharmaceutical quality is essential for manufacturers that want to serve regulated markets. Good Manufacturing Practice (GMP) guidelines establish the standards that help ensure medicines are consistently produced, controlled, and suitable for their intended use. In the European Union, compliance with applicable EU GMP requirements is an important part of pharmaceutical manufacturing and quality assurance.
However, meeting these requirements involves more than passing an inspection. Manufacturers must maintain accurate documentation, validate critical processes, control contamination risks, qualify suppliers, and ensure that employees follow approved procedures. Even small gaps in these areas can lead to inspection findings, production delays, or difficulties in maintaining regulatory compliance.
For pharmaceutical companies, distributors, importers, and brand owners, understanding these challenges is valuable when selecting a manufacturing partner. Working with a European GMP Certified Pharmaceutical Company requires careful verification of the manufacturer's regulatory status, quality systems, and product-specific capabilities.
Medwise Overseas recognizes the importance of evaluating pharmaceutical manufacturing partners through a quality-focused and transparent approach. Understanding common compliance challenges can help businesses ask the right questions and make better-informed sourcing decisions.
What Is EU GMP Compliance in Pharmaceutical Manufacturing?
EU Good Manufacturing Practice refers to the standards and principles that govern the manufacture and quality control of medicinal products within the European regulatory framework. These requirements cover manufacturing operations, personnel, facilities, equipment, documentation, quality control, and other activities that may affect product quality.
EU GMP compliance is intended to help manufacturers consistently produce medicines that meet established quality requirements. Depending on the product and manufacturing activity, relevant requirements may also involve qualification, validation, computerized systems, supplier oversight, and data integrity.
It is important to understand that GMP compliance is not simply a certificate displayed by a company. Buyers should verify the relevant manufacturing authorization, GMP inspection status, product category, and scope of approved activities where applicable.
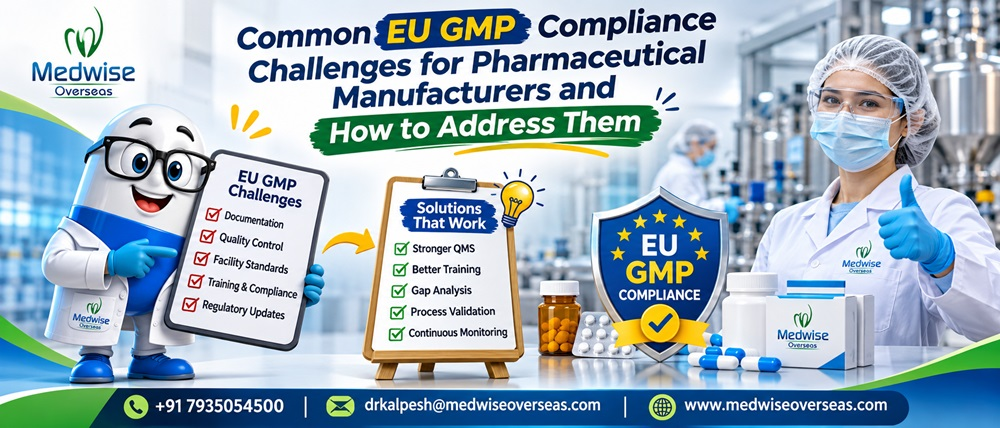
Common EU GMP Compliance Challenges and Their Solutions
1. Incomplete Documentation and Poor Data Integrity
Documentation is one of the foundations of pharmaceutical quality management. Manufacturing records, laboratory results, equipment logs, deviation reports, and batch records must accurately reflect the work performed.
Missing entries, inconsistent records, uncontrolled document versions, or incomplete audit trails can make it difficult to demonstrate that procedures were followed correctly.
How to address this challenge:
-
Maintain controlled and approved standard operating procedures (SOPs).
-
Review batch manufacturing and packaging records for completeness.
-
Establish clear procedures for correcting documentation errors.
-
Apply appropriate access controls and audit-trail reviews to electronic records.
-
Train employees on data integrity principles, including accurate, attributable, and contemporaneous recording.
Regular internal audits can help identify documentation gaps before they become more serious compliance concerns.
2. Inadequate Quality Risk Management
Pharmaceutical manufacturers must identify and assess risks that could affect product quality and patient safety. Risks may arise from raw materials, manufacturing processes, equipment failures, environmental conditions, or changes to established procedures.
A risk assessment that is incomplete or not updated when circumstances change may leave important quality concerns unaddressed.
How to address this challenge:
Manufacturers should apply a structured, science-based risk management process. Tools such as Failure Mode and Effects Analysis (FMEA) can help teams identify potential failures, assess their impact, and prioritize appropriate controls.
Risk assessments should be reviewed when there are significant process changes, recurring deviations, supplier problems, or new information about product quality. Decisions should be documented, justified, and followed by appropriate monitoring.
3. Contamination Control and Cross-Contamination Risks
Contamination can compromise medicine quality and create risks for patients. Depending on the product and manufacturing environment, concerns may include microbial contamination, particulate matter, chemical residues, and cross-contamination between products.
These risks are especially important when facilities manufacture multiple products or handle materials that require specific containment measures.
How to address this challenge:
Manufacturers should develop contamination control strategies appropriate to their products, processes, and facilities. Relevant measures may include equipment cleaning validation, environmental monitoring, personnel hygiene, material segregation, and suitable ventilation or containment systems.
Cleaning procedures should be supported by scientific justification and documented evidence. Where applicable, manufacturers should also assess whether dedicated equipment, dedicated facilities, or additional controls are necessary.
4. Weak Supplier Qualification and Raw Material Control
The quality of a finished medicine depends partly on the quality and consistency of its starting materials, active pharmaceutical ingredients, and packaging components.
Working with suppliers without adequate qualification can introduce risks such as inconsistent material quality, incomplete documentation, supply interruptions, or difficulties tracing the source of materials.
How to address this challenge:
-
Establish risk-based supplier qualification procedures.
-
Verify relevant supplier licenses, quality documentation, and audit information.
-
Define material specifications and acceptance criteria.
-
Review certificates of analysis and perform appropriate incoming-material testing.
-
Monitor supplier performance and investigate recurring quality issues.
Manufacturers should also maintain suitable traceability records and have procedures for responding to material defects or suspected quality problems.
5. Process Validation and Equipment Qualification Gaps
Manufacturers must demonstrate that relevant equipment, utilities, and processes are suitable for their intended use. Insufficient qualification or validation can make it difficult to establish that a manufacturing process consistently delivers the required quality.
Changes to equipment, production parameters, software, or manufacturing methods may also affect the validated state.
How to address this challenge:
A planned lifecycle approach to qualification and validation helps maintain control. Manufacturers should establish documented protocols, predefined acceptance criteria, and appropriate review of results.
Where applicable, process validation, cleaning validation, analytical method validation, and computerized system validation should be addressed according to regulatory requirements and risk. Periodic reviews and change control can help determine whether requalification or revalidation is necessary.
6. Insufficient Employee Training and GMP Awareness
Even well-designed procedures can fail when employees do not understand their responsibilities. Inadequate training may lead to incorrect documentation, procedural deviations, poor hygiene practices, or inconsistent handling of materials.
How to address this challenge:
Manufacturers should provide role-specific GMP training before employees perform relevant activities. Refresher training should be provided according to identified needs, changes in procedures, and applicable requirements.
Training effectiveness can be assessed through practical demonstrations, knowledge checks, supervised activities, and reviews of recurring errors. Training records should clearly show what was covered and whether employees were qualified for their assigned responsibilities.
7. Ineffective Deviation Management and CAPA
Deviations are events in which an activity or result does not follow an approved procedure or expected requirement. A common challenge is correcting the immediate problem without adequately investigating its underlying cause.
Corrective and Preventive Action (CAPA) systems are intended to address identified problems and help prevent recurrence.
How to address this challenge:
Manufacturers should document deviations promptly, assess their potential impact on product quality, and investigate them to an appropriate level. Root-cause analysis should be evidence-based rather than relying on assumptions.
CAPA actions should have assigned responsibilities, defined timelines, and documented effectiveness checks. Reviewing trends across deviations, complaints, and audit findings can help identify recurring weaknesses that require broader improvements.
8. Managing Regulatory Changes and Inspection Readiness
Pharmaceutical regulations and guidance can evolve. Manufacturers must understand which requirements apply to their products, activities, and target markets, while ensuring that relevant changes are implemented in their quality systems.
Poor inspection preparation can expose unresolved documentation gaps, overdue actions, or inconsistencies between written procedures and actual practices.
How to address this challenge:
Companies should monitor relevant regulatory updates, assess their impact, and document implementation decisions. Internal audits, mock inspections, management reviews, and timely closure of corrective actions can support inspection readiness.
Inspection preparation should reflect normal operating practices. Employees must understand their responsibilities and be able to explain the procedures they follow without relying on rehearsed answers.
How to Evaluate a European GMP Certified Pharmaceutical Company
Selecting a manufacturing partner requires more than comparing prices and delivery timelines. Pharmaceutical buyers should evaluate regulatory evidence, quality practices, product suitability, and communication standards.
Before entering a manufacturing agreement, consider the following checks:
-
Regulatory status: Verify applicable manufacturing authorization and GMP inspection evidence through relevant official sources.
-
Product and dosage-form capability: Confirm that the facility is authorized and equipped for the specific product category and manufacturing activity.
-
Quality documentation: Discuss applicable specifications, batch records, testing arrangements, deviation handling, and change-control procedures.
-
Supplier and material controls: Understand how starting materials and packaging components are sourced, qualified, and assessed.
-
Quality agreements: Define responsibilities for complaints, product recalls, deviations, changes, documentation, and other relevant quality activities.
-
Supply reliability: Review realistic production capacity, lead times, communication processes, and contingency planning.
A company should not be considered EU GMP compliant solely because its website uses terms such as GMP certified or European quality. The evidence must relate to the actual facility, activities, and products under consideration.
How Medwise Overseas Can Support Better Pharmaceutical Sourcing Decisions
Medwise Overseas can be considered by businesses exploring pharmaceutical sourcing and manufacturing opportunities. Before proceeding with any proposed arrangement, buyers should discuss the required product range, dosage forms, target markets, documentation, and applicable regulatory expectations directly with the company.
A practical evaluation process can help both parties establish whether the proposed arrangement fits the buyer's commercial and quality requirements.
Potential discussion points include:
-
Product availability and manufacturing or sourcing arrangements.
-
Applicable licenses, quality documents, and regulatory evidence.
-
Product specifications, packaging requirements, and documentation needs.
-
Expected order quantities, delivery timelines, and supply planning.
-
Procedures for handling quality concerns and changes, where relevant.
Specific certifications, manufacturing capabilities, export permissions, and product approvals should be confirmed with Medwise Overseas before being presented as established company credentials.
Conclusion
EU GMP compliance requires ongoing attention to documentation, contamination control, supplier qualification, validation, employee training, and effective quality management. Addressing these challenges through risk-based procedures, regular reviews, and documented corrective actions helps pharmaceutical manufacturers maintain control over their operations.
For pharmaceutical buyers, the same principles provide a practical framework for evaluating potential suppliers and manufacturing partners. Rather than relying only on marketing claims, businesses should verify regulatory evidence, assess product-specific capabilities, and clarify quality responsibilities before placing an order.
Medwise Overseas encourages prospective partners to discuss their product requirements, documentation expectations, and sourcing objectives directly. A transparent evaluation process can help establish a stronger foundation for responsible pharmaceutical sourcing and long-term business relationships.
1. What is EU GMP compliance?
Medwise Overseas Pvt. Ltd. offers end-to-end pharmaceutical manufacturing and global export solutions. Our product range includes tablets, capsules, syrups, injectables, and healthcare products. We specialize in customized manufacturing, dossier preparation, product marketing, and technology transfer, supporting compliant product development and international supply. Visit Website : https://www.medwiseoverseas.com/
Comments (0)